Division of Nephrology
Sanna-Cherchi Lab
ENVISIONING SCIENCE-BASED PRECISION MEDICINE THROUGH GENE DISCOVERY AND FUNCTIONAL MODELING
Understanding the Basis of Recurrent Focal Segmental Glomerulosclerosis
Recurrent focal segmental glomerulosclerosis (rFSGS) is a rare and understudied glomerular disease that presents significant clinical challenges. It is characterized by a primary podocytopathy with histologic findings of FSGS that recurs, usually within few weeks, after kidney transplant often leading to rapid graft loss. Its rarity has led to limited ability in designing studies to address its pathophysiology, inadequate treatments, and poor patient outcomes. Advances in FSGS genetics have mostly focused on Mendelian forms of FSGS, which, differently from rFSFS, do not recur post-transplant, leaving the genetic basis and pathobiology of rFSGS unresolved and hindering effective risk stratification and targeted therapy. To address these gaps, we have established a coalition of investigators across the world to enroll, clinically characterize, and biosample a uniquely large cohort of individuals affected by rFSGS. This coalition leverages on both established cohorts of thousands of cases with idiopathic nephrotic syndrome, and on prospective biobanking for immunogenetic and multiomic studies, with the ultimate goals of enhancing understanding of rFSGS genetics and mechanisms of disease, and improving diagnostic and therapeutic strategies.
Investigators: Elena Martinelli, MD; Tze Yin (Joanne) Lim, MS; Juntao Ke, PhD
Project Coordination: Massiel Baez Belen

Barry, et al., “Multi-population genome-wide association study implicates immune and non-immune factors in pediatric steroid-sensitive nephrotic syndrome.” Nat Commun. 2023 Apr 29;14(1):2481
Genome-wide Association Study (GWAS) to understand the complex etiology of Nephrotic Syndrome
Focal Segmental Glomerulosclerosis (FSGS) is a major contributor to end-stage renal disease, and presents a significant global health challenge with limited treatment options and an incomplete understanding of its underlying biology. To enhance our understanding of the genetic and molecular etiology of this disease, and others that lead to nephrotic syndrome (NS), we integrate cutting-edge human and mouse genetics alongside the analysis of bulk, single-cell and “multiome” transcriptomics. Our genome-wide association studies (GWAS) leverage a substantial cohort of 5,600 meticulously characterized FSGS patients, spanning diverse demographics and genetic backgrounds, where we have uncovered associations with known FSGS-associated genes, including APOL1, and with novel candidates, including those on Chromosomes 4 and 15. These findings hold promise for advancing our understanding of FSGS pathogenesis and improving targeted treatments for affected individuals.
Investigators: Yask Gupta, PhD; Juntao Ke, PhD; Tze Yin (Joanne) Lim, MS
Genetic modifiers of APOL1-associated FSGS
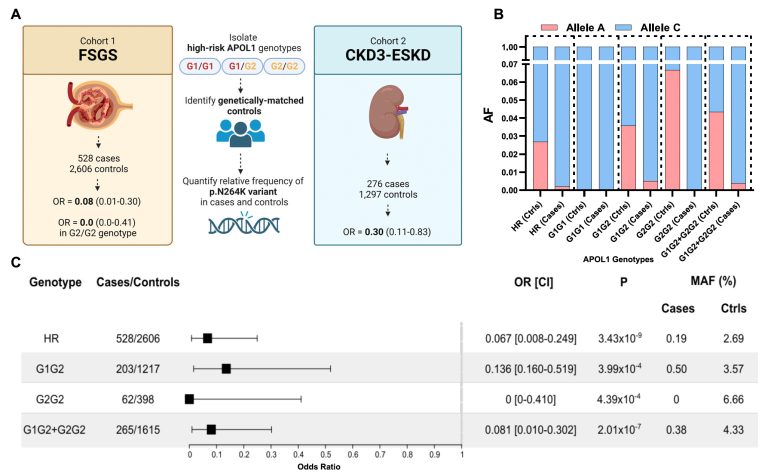
In this project, we are investigating the modifiers for APOL1 high-risk genotypes in FSGS, addressing the relatively lower FSGS occurrence in African populations. Utilizing advanced statistical methods and diverse datasets, our findings to-date interestingly highlight the distinctiveness of the G2-p.N264K haplotype compared to the common G2 risk allele. By analyzing APOL1-HR FSGS subjects and healthy APOL1-HR controls, we reveal that the p.N264K minor allele ‘A’ provides robust protection, particularly in G2 allele carriers (G1/G2 and G2/G2). Notably, this allele is entirely absent in G1/G1 subjects, ensuring complete FSGS protection in G2/G2 cases. Additionally, G1/G2 individuals exhibit significant protection, and combining G1/G2 and G2/G2 cases strengthens these findings. In summary, our results underscore the substantial protective effect of the APOL1 p.N264K missense variant against APOL1-associated FSGS, primarily within G2-containing APOL1 high-risk genotypes of African origin. Practically, based on our analysis, carrying one copy of the p.N264K missense variant reduces the risk of developing FSGS by at least 8.3 times, offering valuable insights into FSGS genetics.
Investigators: Yask Gupta, PhD; Juntao Ke, PhD; Tze Yin (Joanne) Lim, MS

Rare Variant Landscape of nephrotic syndrome caused by FSGS and MCD

Weng et al., “De novo TRIM8 variants impair its protein localization to nuclear bodies and cause developmental delay, epilepsy, epilepsy, and focal segmental glomerulosclerosis.” Am J Hum Genet. 2021 Feb 4;108(2):357-367
TRIM8 Syndrome and Focal Segmental glomerulosclerosis (FSGS)
Working with our collaborators, in a large cohort of FSGS/SRNS patients we identified a novel set of truncating variants in TRIM8 that underlie a neuro-renal syndrome in children, characterized by developmental delay, epilepsy, and FSGS. To test our hypothesis that de novo heterozygous truncating mutations in TRIM8 are the cause of a novel, clinically recognizable syndrome characterized by early-onset epilepsy and nephrotic syndrome caused by FSGS, we are employing animal models, in vitro modeling using human/patient-derived iPSCs to produce kidney organoids, transcriptomic analyses at the single cell level, and deep cellular phenotyping.
Investigators: Jeremiah Martino, PhD; Qingxue Liu, MS
Collaborators: Erica Davis, PhD; Patty Weng, MD; Rosemary V. Sampogna, MD, PhD; Dietrich Egli, PhD
For questions on this page’s content(s), please contact webmaster